By Ramesh Palav | Pharma Manufacturing Hub
Pharmaceutical packaging is the final operational stage before a finished medicine reaches the patient, but it is far more than placing tablets or capsules into a blister, bottle, or carton. Every packaging component, printed detail, sealing operation, and reconciliation record must be controlled to prevent product mix-ups, labeling errors, contamination, and traceability failures.
In an oral solid dosage (OSD) manufacturing facility, effective quality checks during pharmaceutical packing operations require close coordination between Production, In-Process Quality Control (IPQC), Quality Assurance (QA), Engineering, and Warehouse teams. A packaging line may run smoothly from a mechanical perspective and still produce unacceptable packs if the wrong foil is loaded, a batch code is incorrect, a leaflet is missing, or a rejection system fails to remove a defective pack.
The purpose of this guide is to explain how these checks should be performed on the shop floor, what personnel should verify, how results should be documented, and what actions are required when a defect is detected. The actual inspection frequencies, acceptance criteria, and responsibilities must always follow approved procedures, product specifications, risk assessments, and applicable regulatory requirements.
Table of Contents
- Understanding pharmaceutical packing operations
- Pre-packaging line clearance
- Packaging material verification
- Quality checks during primary packaging
- Quality checks during secondary packaging
- Sampling frequency and documentation
- Packaging material reconciliation and yield verification
- GMP documentation and data integrity
- Handling packaging defects and deviations
- Roles and responsibilities
- Practical shop-floor case study
- Pharmaceutical packing quality checklist
- Regulatory requirements
- Frequently asked questions
- Conclusion
1. Understanding Pharmaceutical Packing Operations
Pharmaceutical packing protects the product, provides essential information to patients and healthcare professionals, and maintains identification and traceability throughout distribution.
For tablets and capsules, packaging operations generally involve two stages.
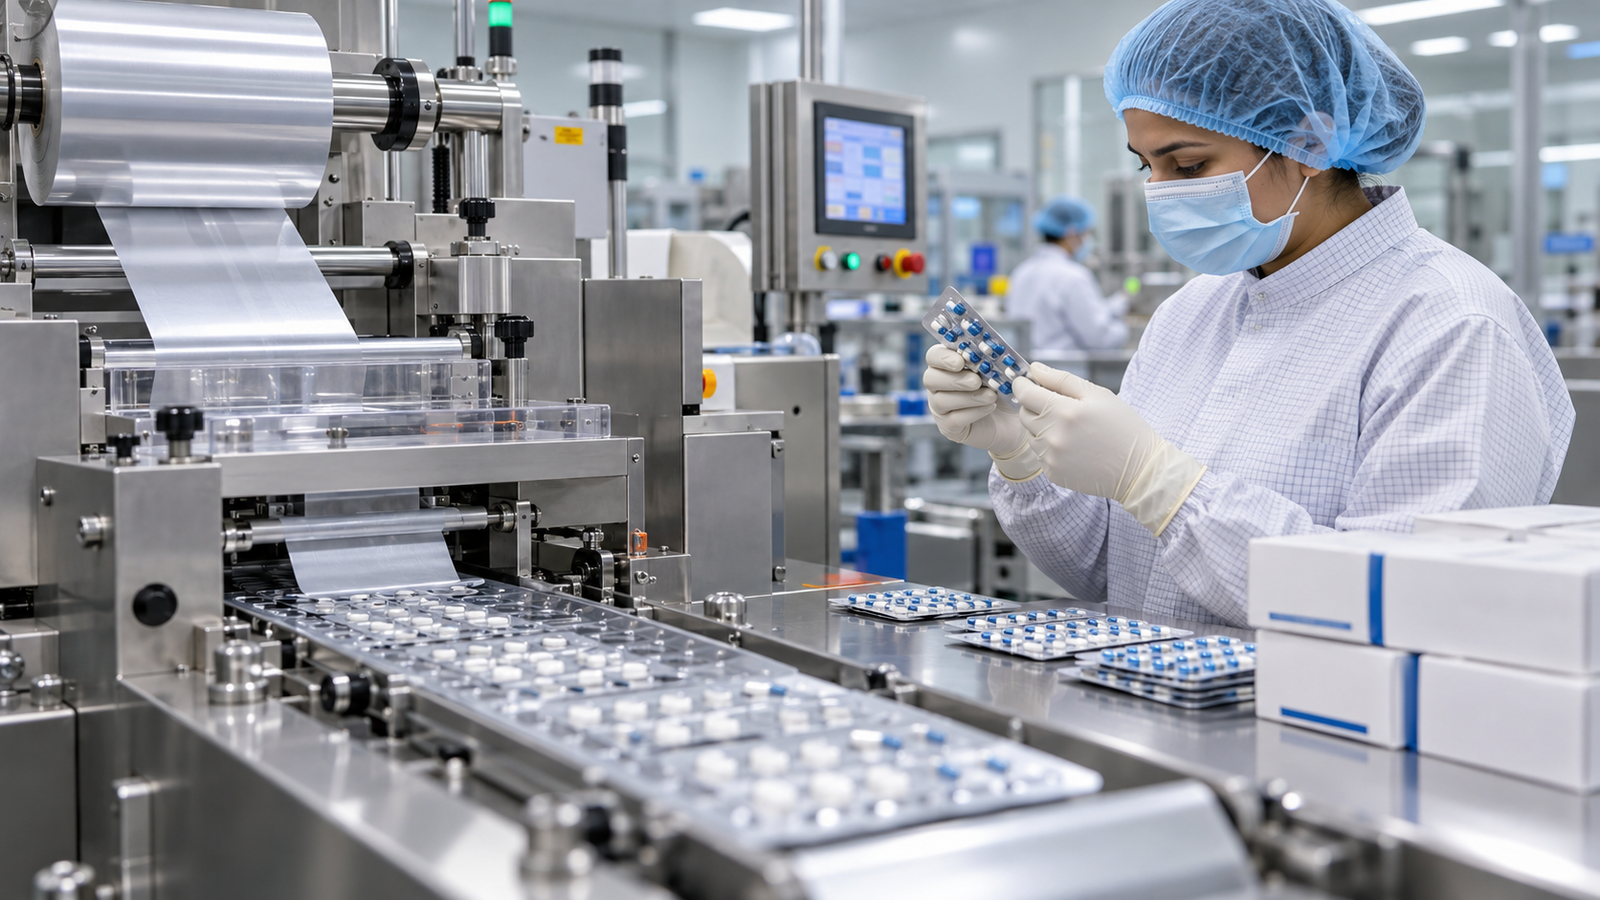
Primary packaging places the product in direct contact with its immediate packaging system or within the product’s immediate container. Typical examples include blister packs, strip packs, and bottles.
Secondary packaging groups and protects primary packs using cartons, package inserts, labels, and shipper boxes. Depending on the product and market, it may also involve serialization, aggregation, and other traceability controls.


10
Quality failures can occur at either stage. A blister may contain a broken tablet, a foil may have a defective seal, a carton may contain the wrong leaflet, or a shipper may carry an incorrect product identification label. Each defect has a different potential impact, but all require an appropriate control and documented response.
The central principle is straightforward: every pack must contain the correct product, in the correct quantity, with the correct packaging components and accurate information.
2. Pre-Packaging Line Clearance
Line clearance is one of the most important GMP checks before starting a packaging batch. Its purpose is to ensure that materials, documents, labels, and products from a previous operation cannot mix with the current batch.
A line clearance should be performed according to the approved pharmaceutical packing SOP and documented before the line is released for use.
2.1 Removal of previous batch materials
The responsible personnel should verify that all previous-batch materials have been removed from the packaging machine, conveyors, accumulation tables, hoppers, coding stations, inspection stations, and surrounding work areas.
Check for:
- Leftover tablets, capsules, blisters, cartons, labels, and leaflets.
- Previous batch documents and loose printed material.
- Obsolete or unused coding details in the printer or coding software.
- Material inside accessible machine guards, chutes, and collection trays.
- Rejected packs and material in designated rejection containers.
- Unidentified containers or packaging components in the line area.
A visual check of the main machine alone is not sufficient. Small quantities of printed material can remain in less visible locations and create a serious mix-up risk.
2.2 Equipment cleaning and status verification
Confirm that the equipment has been cleaned as required and that its cleaning and operational status are acceptable. Check the relevant equipment identification, cleaning records, maintenance status, and any required pre-use checks.
If the previous product or batch has a specific cleaning or segregation requirement, the applicable procedure must be followed before the next batch is introduced.
2.3 Verification of the current batch
Before releasing the line, verify the current batch’s:
- Product name and strength.
- Dosage form and batch number.
- Approved batch packaging instructions.
- Packaging configuration and market presentation.
- Approved packaging material specifications.
- Required coding details and packaging order.
The details on the packaging order, batch packaging record, material labels, and approved coding instructions must agree.
2.4 Line clearance documentation
The person performing the clearance should complete the required checklist, record observations, and obtain the required independent verification or QA authorization where specified by the site’s procedures.
Practical point: Do not treat line clearance as a signature exercise. The check must be physically performed, and any discrepancy must be resolved before the line is released.
3. Verification of Packaging Materials
Packaging material verification confirms that only approved, correct, and suitable components are used for the intended product and batch. This check is particularly important when several products, strengths, or pack sizes are handled in the same manufacturing facility.
3.1 Verify material identity and approval status
Before loading a packaging material onto the line, compare its identification label with the approved packaging instructions and material specifications.
Verify the following:
| Packaging material | Important checks |
|---|---|
| Blister forming film | Material code, specification, width, thickness where required, appearance, and suitability |
| Printed lidding foil | Product identity, strength, artwork, print details, and material code |
| Strip foil | Correct material, approved print, dimensions, and appearance |
| Cartons | Product name, strength, pack size, artwork version, and required statutory information |
| Package inserts | Correct product, language, approved version, and legibility |
| Labels | Product identity, strength, presentation, and applicable coding information |
| Shipper cartons | Correct product, batch identification requirements, quantity, and shipping details |
The checks should include material status, such as approved or released for use, as required by the site’s material management system.
3.2 Verify printed packaging components
Printed components require particular attention because they carry information that identifies the medicine and communicates essential instructions.
Check the approved artwork or reference sample for:
- Product name and strength.
- Dosage form and pack configuration.
- Manufacturer or marketer details, as applicable.
- Batch number and expiry information, where printed.
- Manufacturing date, MRP, and other legally required details for the intended Indian market.
- Required warnings, storage conditions, and statutory declarations.
- Barcode, QR code, or serialization information where applicable.
The exact content depends on the product, approved labeling, destination market, and current legal requirements.
3.3 Check material condition before use
Packaging materials should be inspected for visible damage, contamination, moisture exposure, deformation, poor printing, missing text, and other defects that may affect their suitability.
For example, a carton may have the correct product name but contain a printing defect that obscures the strength. The material should not be accepted simply because the product name is readable.
Similarly, damaged forming film or lidding foil can cause sealing problems, poor appearance, or compromised pack integrity.
3.4 Verify coding and barcode systems
Where applicable, ensure that the correct coding recipe, printer settings, barcode configuration, and product master data are selected.
The operator should verify the printed output against the approved reference rather than relying only on the machine display.
For barcode and serialization systems, perform the required verification using approved equipment and procedures. A code that looks visually correct may still fail scanning or contain incorrect encoded information.
Practical point: Verification of printed material before use and verification of the actual printed output during the run are two separate controls. Both are necessary.
4. Quality Checks During Primary Packaging
Primary packaging checks depend on the packaging technology and product. Blister and strip packing operations require particular attention to product placement, forming, sealing, coding, and pack integrity.


10
4.1 Blister appearance and pocket formation
Inspect the blister for uniform pocket formation, acceptable appearance, correct dimensions, and absence of defects.
Look for:
- Incomplete or distorted pockets.
- Cracks, pinholes, tears, or other visible damage.
- Poor forming or uneven pocket depth.
- Misalignment between formed cavities and lidding foil.
- Excessive deformation or damage during cutting.
- Burrs, sharp edges, or irregular perforations where relevant.
The acceptance criteria must come from the approved specification and validated or otherwise appropriately qualified process requirements.
4.2 Tablet and capsule placement
Verify that every pocket contains the correct dosage unit and that the product is properly positioned.
Inspect for:
- Empty pockets.
- Two tablets or capsules in one pocket when only one is intended.
- Broken, chipped, cracked, or visibly damaged dosage units.
- Wrong product or strength.
- Improperly positioned tablets or capsules that may interfere with sealing.
- Foreign particles or other visible contamination.
The number of units per blister and the cavity arrangement must match the approved pack configuration.
For capsules, check for visible shell damage, separation, leakage of contents, or other defects relevant to the product.
4.3 Sealing quality and integrity
Sealing quality is a critical control because the blister or strip must protect the product throughout its intended shelf life.
Check for:
- Wrinkles or folds in the sealing foil.
- Incomplete sealing or visible unsealed areas.
- Burn marks, excessive discoloration, or heat damage.
- Foil misalignment.
- Contamination or trapped product particles in the sealing area.
- Evidence of poor adhesion or other visible sealing defects.
Where required, perform a leak test or another approved integrity test using the specified method and equipment.
Do not rely on appearance alone when the process requires a separate integrity test. A blister may look acceptable but still fail a validated leak-detection method.
4.4 Batch coding and overprinting
Inspect the actual code printed on the blister or strip.
Verify:
- Product and batch identification, where required.
- Manufacturing date and expiry date.
- Legibility, contrast, alignment, and permanence as specified.
- Correct coding format and placement.
- Consistency with the approved batch packaging record and coding instructions.
At start-up, the first acceptable printed output should be checked and approved by the designated personnel according to the site’s procedure. Further checks must follow the established inspection plan.
After a printer adjustment, material change, or other relevant intervention, verify the output again as required by the approved SOP.
4.5 Foil alignment, cutting, and perforation
Check that the blister or strip is cut to the correct dimensions and that perforations, if present, are properly formed.
Defects may include:
- Incorrect pack dimensions.
- Incomplete cuts or excessive burrs.
- Misaligned perforations.
- Damaged sealing areas.
- Inconsistent separation between individual packs.
- Incorrect registration between printed foil and the formed blister.
These defects can affect pack usability, presentation, and, in some cases, package integrity.
4.6 Rejection system verification
Automated inspection and rejection systems can help prevent defective packs from progressing to secondary packaging. Their effectiveness depends on correct setup, challenge testing, and control of rejected material.
Where applicable, verify the detection and rejection system using approved challenge procedures for relevant defects, such as missing product, incorrect coding, or other configured inspection failures.
Confirm that:
- The challenge is performed using the approved method.
- The system detects the intended defect.
- The defective pack is rejected into the designated collection point.
- Rejected packs are not inadvertently returned to the good-product stream.
- Rejection counters or electronic records are checked as required.
- Challenge results are documented.
Challenge testing should be performed at the frequency and under the conditions specified by the approved procedure. Do not assume that every machine uses the same challenge frequency or detection capability.
5. Quality Checks During Secondary Packaging
Secondary packaging introduces additional risks because multiple components must be assembled into the correct finished pack.
A blister may pass every primary packaging check, yet the finished product can still be unacceptable if the carton contains the wrong leaflet or carries incorrect labeling.
5.1 Correct carton and product combination
Verify that the carton matches the product, strength, market, pack size, and approved artwork.
Check that the correct number of blisters or strips is inserted into each carton. For bottle packaging, verify the specified bottle, closure, seal, label, and quantity of tablets or capsules, as applicable.
The packaging configuration must match the approved batch packaging instructions.
5.2 Package insert or leaflet verification
Check that the correct package insert is present, belongs to the correct product and market, and is inserted in the required manner.
Common defects include:
- Missing leaflet.
- Incorrect product leaflet.
- Wrong language version.
- Folded, torn, or illegible leaflet.
- Multiple leaflets where only one is required.
Where automated leaflet detection is installed, verify its functionality as required by the approved procedure. Do not assume that the presence of a detection device eliminates the need for the prescribed in-process checks.
5.3 Carton coding and labeling
Inspect the carton for correct and legible printing.
Confirm applicable details, including:
- Product name and strength.
- Batch number.
- Manufacturing and expiry dates.
- MRP and other statutory declarations for the intended market.
- Required warnings and storage instructions.
- Barcode or serialization information, where applicable.
Compare the actual output with the approved coding instructions and reference. Verify both the printed characters and their correct association with the batch.
5.4 Carton closure and appearance
Check the finished carton for correct folding, closure, and overall appearance.
Look for:
- Open or partially closed cartons.
- Poor glue application or inadequate adhesion.
- Damaged or crushed corners.
- Torn carton flaps.
- Misaligned labels.
- Excessive adhesive or visible contamination.
- Cartons with unreadable or damaged printing.
Where tamper-evident features are required, inspect them according to the approved specifications.
5.5 Barcode, serialization, and aggregation
For markets or products requiring serialization, confirm that the applicable identification codes are correct and readable, and that the system records the required events.
Where aggregation is implemented, verify the relationship between individual saleable packs, cases, and pallets according to the approved system and market requirements.
Potential failures include duplicate codes, unreadable codes, incorrect parent-child relationships, failed commissioning, and discrepancies between physical packs and electronic records.
Affected packs should be handled under the applicable procedure. A failed scan or serialization event must not be bypassed simply to maintain production speed.
5.6 Shipper carton and pallet checks
Before transferring finished cartons to the next stage, verify:
- Correct number of saleable packs per shipper.
- Correct product and batch identification.
- Appropriate shipper labels.
- Correct quantity and packaging configuration.
- Required pallet identification and traceability.
- Legibility and consistency of shipping information.
The final shipper should be traceable to the relevant product and batch records.
6. Sampling Frequency and Documentation
Quality checks are effective only when they are performed at suitable intervals and the results are documented accurately.
There is no single sampling frequency that is appropriate for every pharmaceutical packing line. The inspection plan should be established through approved procedures, applicable regulatory requirements, process knowledge, product risk, equipment capability, and relevant validation or qualification evidence.
6.1 When should checks be performed?
The approved inspection plan may require checks at the following stages:
| Stage | Examples of checks |
|---|---|
| Before start-up | Line clearance, material identity, coding setup, equipment status |
| Start-up | Initial printed output, product placement, sealing, pack configuration |
| During routine operation | Coding, pack appearance, completeness, sealing and other specified attributes |
| After a material or batch change | Correct material identity, new coding details, applicable line clearance and verification |
| After machine adjustment | Recheck affected attributes and verify acceptable output |
| After stoppage or breakdown | Assess affected packs and perform checks required by the procedure |
| At line closure | Final reconciliation, material return or destruction, batch documentation and line clearance |
These are general examples, not a universal mandatory sampling schedule. The approved SOP must specify the actual frequency, sample size, acceptance criteria, and responsibilities.
6.2 How to perform an in-process check
A practical sequence is:
- Collect the sample according to the approved sampling procedure.
- Identify the batch, line, time, and relevant process stage.
- Compare the sample against approved specifications, reference samples, or coding instructions.
- Record each required result, including failures.
- If a result is unacceptable, take the action defined by the deviation and nonconformance procedure.
- Verify that required corrective actions are completed and obtain authorization before restarting, where applicable.
Sampling must represent the intended process and should not be selected merely because the packs look good.
6.3 Record results promptly
Record the date, time, result, operator or inspector identification, and any other required information in the Batch Packaging Record (BPR), in-process inspection sheet, or validated electronic system.
If a result is outside the acceptance criteria, document the actual observation and follow the applicable procedure. Do not replace a failed result with a passing result from a later sample without retaining and assessing the original failure.
7. Packaging Material Reconciliation and Yield Verification
Reconciliation is an important control for identifying unexplained losses, preventing reuse of unauthorized printed components, and detecting possible packaging mix-ups.
7.1 Reconcile issued and used materials
Compare the quantity issued with the quantity consumed, returned, rejected, sampled, damaged, or otherwise accounted for.
For printed components, reconciliation may include:
- Cartons.
- Labels.
- Leaflets.
- Printed foil.
- Shipper labels and other batch-specific material.
The precise reconciliation method depends on the component, the packaging process, and the approved procedure.
7.2 Investigate discrepancies
Suppose a packaging batch was issued 10,000 printed cartons, but the documented quantities account for only 9,850 cartons. The remaining 150 cartons must be accounted for according to the approved reconciliation method.
The team should check recorded usage, rejects, samples, returns, and other authorized dispositions. The example is illustrative; actual limits and reconciliation rules must come from the relevant procedure.
If the discrepancy exceeds an established limit, or indicates a possible mix-up or loss of printed material, escalate it for documented investigation and QA assessment.
Never adjust reconciliation figures simply to make the quantities balance. Any correction must be supported by evidence and follow the approved documentation and investigation process.
7.3 Finished product yield
Compare actual finished-pack quantities with the expected quantity, taking account of the approved calculation method and documented losses.
Unexpected yield differences can indicate counting errors, unrecorded rejects, product loss, or a possible discrepancy in the packaging process. These should be evaluated rather than automatically attributed to normal wastage.
8. GMP Documentation and Data Integrity
Documentation provides evidence that the packaging process was performed according to approved instructions and that required quality checks were completed.
A well-maintained batch packaging record should allow a reviewer to reconstruct what happened during the operation, which materials were used, what checks were performed, what defects occurred, and how discrepancies were resolved.
8.1 Good Documentation Practices
Follow these principles when recording packing activities:
- Record information at the time the activity is performed.
- Use the approved forms, logbooks, and electronic systems.
- Record actual results rather than expected results.
- Identify the person performing the check as required.
- Document deviations, equipment stoppages, adjustments, and other reportable events.
- Make corrections according to the applicable procedure, preserving the original entry and recording the required reason and attribution.
- Ensure records remain complete, readable, retrievable, and protected against unauthorized changes.
Never record an inspection as completed if it was not performed.
8.2 Applying ALCOA+ principles
ALCOA+ is a useful framework for maintaining reliable GMP records.
| Principle | Practical application during packing |
|---|---|
| Attributable | The record identifies who performed or reviewed the activity |
| Legible | Results can be read and understood |
| Contemporaneous | Entries are made when the activity occurs |
| Original | The original record or an appropriate verified copy is retained |
| Accurate | Recorded information reflects the actual result |
| Complete | Required results, failures, and relevant events are included |
| Consistent | Entries follow a logical sequence with appropriate dates and times |
| Enduring | Records remain preserved for the required retention period |
| Available | Records can be retrieved for review, investigation, or inspection |
8.3 Control of electronic systems
Where electronic batch records, vision inspection systems, printers, or serialization platforms are used, the site should apply the controls required by the applicable regulations and approved system procedures.
Depending on the system and intended use, these may include:
- Authorized user access and appropriate permissions.
- Validated functionality for the intended use.
- Audit trails and review of relevant changes.
- Secure retention and retrieval of electronic records.
- Control of master data, recipes, and coding parameters.
- Backup and recovery arrangements.
- Appropriate management of system changes.
21 CFR Part 11 applies to electronic records and signatures within its scope; it does not automatically apply to every electronic device on a packaging line. Applicability must be assessed against the relevant regulatory requirements and system use.
9. Handling Packaging Defects and Nonconformities
When a packaging defect is identified, the objective is to prevent potentially affected product from progressing, establish the extent of the problem, and ensure that the final disposition is supported by evidence.
The response should be proportionate to the defect and follow the site’s approved deviation, nonconformance, and product-disposition procedures.
9.1 Immediate actions
For a defect that could affect product quality, labeling correctness, patient safety, or traceability:
- Stop the affected operation when warranted by the risk and approved procedure.
- Identify and segregate potentially affected packs and materials.
- Establish the relevant batch, line, time period, and last known acceptable check.
- Inform the responsible Production supervisor and QA.
- Record the observation and preserve relevant evidence.
- Prevent release or further processing of affected material until the required assessment is completed.
Do not assume that only the pack in which the defect was found is affected. The investigation must establish a justified scope.
9.2 Common defects and appropriate responses
| Defect | Key response |
|---|---|
| Incorrect batch or expiry code | Stop or hold affected output, segregate packs, and assess the period since the last acceptable verification |
| Missing tablet or capsule | Check the product feeding and inspection system, identify affected output, and investigate the cause |
| Missing or incorrect leaflet | Hold affected packs and verify leaflet supply, insertion, and detection controls |
| Poor blister sealing | Assess seal-related process parameters, integrity results, and the potentially affected quantity |
| Incorrect printed foil or carton | Prevent further use, segregate affected materials and packs, and investigate possible mix-up exposure |
| Barcode or serialization failure | Hold affected packs as required and resolve the failure through the approved system procedure |
| Reconciliation discrepancy | Secure remaining printed components, check all accounted quantities, and initiate investigation as required |
9.3 Root cause analysis and CAPA
An investigation should identify why the defect occurred and whether similar failures could exist elsewhere.
Depending on the circumstances, possible causes may include:
- Incorrect material issued to the line.
- Inadequate line clearance.
- Incorrect printer recipe or master data.
- Machine adjustment or setup error.
- Worn or damaged tooling.
- Inadequate operator training.
- Failure of a detection or rejection mechanism.
- Ineffective procedural controls.
Root cause analysis should be evidence-based rather than assigning blame without investigation. CAPA should address the identified cause and be proportionate to the risk. Effectiveness should be verified where required by the quality system.
9.4 Restart and disposition
Production should resume only after the required checks and corrective actions have been completed and the designated authority has authorized restart in accordance with site procedures.
QA should assess the quality impact and determine or approve the appropriate disposition within its assigned authority. Rework or repacking must not be performed informally; it requires an approved, documented approach and appropriate assessment.
10. Roles and Responsibilities
Effective packing quality depends on clear responsibilities and coordination between functions.
| Function | Main responsibilities |
|---|---|
| Packing operators | Perform assigned checks, operate equipment according to instructions, identify defects, and record results |
| Production officers and supervisors | Ensure line readiness, verify operational controls, monitor performance, and respond to abnormalities |
| IPQC, where assigned | Perform or review designated in-process inspections and document findings |
| Quality Assurance | Provide quality oversight, review deviations, assess impact, and authorize decisions within its defined authority |
| Engineering and maintenance | Maintain equipment condition, troubleshoot failures, and support approved repairs and adjustments |
| Warehouse and packaging-material personnel | Supply correct approved materials, maintain identification and status controls, and support reconciliation and returns |
The precise allocation of duties may differ between facilities. It must be defined in the approved quality system, SOPs, job responsibilities, and batch packaging instructions.
11. Practical Shop-Floor Case Study: Incorrect Expiry Date on Blister Foil
Consider a blister packing line running a tablet batch. During a routine in-process check, an operator notices that the expiry date printed on the blister foil does not match the approved batch packaging instructions.
The line has been operating for some time, and several blister strips have already passed the coding station.
Step 1: Stop and contain
The operator immediately informs the supervisor and stops the affected operation in accordance with the site’s procedure. Potentially affected packs are identified and segregated so that they cannot be mixed with acceptable output.
Step 2: Establish the affected period
The team identifies the last documented acceptable coding check and determines when the incorrect code may have started appearing.
Packs produced during the potentially affected period are held pending assessment. If the last acceptable check cannot establish a reliable boundary, the investigation should account for that uncertainty.
Step 3: Investigate the coding setup
The team reviews the printer recipe, batch packaging instructions, coding setup records, operator actions, and any recent printer adjustments or interruptions.
The investigation determines whether the error arose from an incorrect recipe, an input error, an inappropriate master-data selection, or another cause. The cause must be supported by evidence.
Step 4: Assess product impact
The team establishes the quantity of potentially affected packs and checks whether any have moved to secondary packaging or another controlled area.
The investigation also considers whether any affected packs have escaped the immediate packaging process and whether additional containment is necessary.
Step 5: Document the event and correct the cause
The discrepancy is recorded through the applicable quality system. The cause is investigated, corrective actions are defined, and relevant checks are repeated using the approved coding instructions.
The affected packs remain segregated until an authorized disposition decision is made.
Step 6: Verify and authorize restart
Before restarting, the responsible personnel verify the correct coding setup and confirm acceptable output according to the approved procedure. The required authorization is documented.
The event may also require CAPA, such as improved recipe verification or additional independent checks, depending on the investigation findings.
Key lesson: Detecting an incorrect expiry date is only the first step. Effective control requires containment, traceability, investigation, documented correction, and a justified decision on the affected product.
This case study is illustrative. The actual response must follow the site’s approved procedures and applicable regulatory requirements.
12. Pharmaceutical Packing Quality Checklist
The following checklist is a practical summary for use when developing or reviewing a site-specific packing checklist. It does not replace the approved SOP or batch packaging record.
Packing line verification checklist
0/28 checked
1. Line readiness
Previous product and batch materials removedEquipment cleaned and status verifiedCorrect product, batch, strength, and pack configuration confirmedLine clearance documented and required authorization obtained
2. Packaging materials
Materials identified and approved for useCorrect artwork, material codes, and specifications verifiedPrinted components and coding instructions checkedMaterial condition acceptable
3. Primary packaging
Correct tablet/capsule placement and pack countBlister or strip appearance acceptableSealing and integrity checks completed as requiredBatch coding and print quality verifiedCutting, perforation, and pack dimensions acceptableRequired detection and rejection challenges passed
4. Secondary packaging
Correct carton and product combinationCorrect number of blisters or strips per cartonCorrect leaflet or insert includedBatch, expiry, MRP, and other required details verifiedBarcode and serialization checks completed where applicableCarton closure and appearance acceptableShipper and pallet identification verified
5. Documentation and closure
In-process checks recorded at the required timesDefects and deviations documented and assessedRejected material accounted for and controlledPackaging material reconciliation completedYield and finished quantity verifiedUnused printed materials returned or destroyed as authorizedFinal documentation review and line closure completed
Reset checklistCopy checklist
For practical planning and training. Complete each item only after the corresponding check has actually been performed. Use approved site records for official GMP documentation.
13. Regulatory Requirements Relevant to Pharmaceutical Packing
Packing controls should be aligned with the regulations applicable to the manufacturing site and the intended market.
WHO GMP
WHO GMP guidance addresses packaging, labeling, documentation, and the prevention of mix-ups and false labeling. It provides a useful foundation for designing and reviewing packaging controls.

World Health Organization
+1
Indian Schedule M
India’s revised Schedule M, notified in December 2023, establishes GMP requirements for pharmaceutical products. Manufacturers should apply the provisions relevant to packaging operations, material control, documentation, and quality management, taking account of applicable implementation requirements and any subsequent amendments.
E-Gazette
US FDA requirements
For products subject to US FDA requirements, 21 CFR Part 211 includes requirements for written packaging and labeling procedures, prevention of mix-ups, material suitability checks, and documented inspection of packaging areas before use. Section 211.130 is particularly relevant to packaging and labeling operations.
GovInfo
EU GMP
EU GMP Volume 4 includes requirements relevant to production, packaging, labeling, documentation, and quality management. Manufacturers supplying the EU should consult the current applicable chapters and annexes.
European Commission
Electronic records and serialization
Electronic record controls, serialization, and traceability requirements depend on the applicable jurisdiction, product, and system. The manufacturer should verify the current legal requirements for each intended market rather than treating one country’s controls as universally applicable.
The regulatory references above provide a starting point for technical review. Before publication or implementation, confirm the latest applicable versions and any product-specific or market-specific requirements.
14. Frequently Asked Questions
1. What are the most important quality checks during pharmaceutical packing operations?
The essential checks include line clearance, packaging material verification, correct product and pack configuration, batch coding, sealing integrity, pack completeness, leaflet inclusion, rejection system performance, reconciliation, and GMP documentation.
The specific checks depend on the packaging technology, product, and applicable requirements.
2. How frequently should in-process packing checks be performed?
The frequency should be defined in the approved SOP and batch packaging instructions, considering product and process risks, regulatory requirements, and equipment capability. Additional checks may be necessary after adjustments, changeovers, breakdowns, or other relevant interventions.
There is no universal inspection interval suitable for every packaging line.
3. Why is line clearance important in pharmaceutical packaging?
Line clearance prevents materials, labels, documents, and products from a previous batch remaining in the work area and mixing with the current batch. It is a key control against product mix-ups and incorrect labeling.
4. What should an operator do if an incorrect batch number or expiry date is detected?
The operator should notify the responsible supervisor, stop the affected operation when warranted, and ensure that potentially affected packs are identified and segregated. The team should establish the affected period, investigate the cause, document the event, and obtain the required authorization before restarting.
5. What is packaging material reconciliation?
Packaging material reconciliation compares quantities issued, used, rejected, returned, destroyed, or otherwise accounted for under the approved procedure. It helps identify unexplained losses and possible mix-ups involving printed packaging materials.
6. What is the difference between IPQC checks and QA oversight during packing?
IPQC personnel perform or review designated in-process inspections according to assigned responsibilities. Production personnel also perform specified operational checks. QA provides quality-system oversight, assesses deviations, and makes or approves quality decisions within its defined authority. The exact allocation varies by site.
7. How does data integrity affect pharmaceutical packing operations?
Data integrity ensures that inspection results, batch coding checks, rejection records, reconciliation figures, and electronic records accurately reflect the work performed. Reliable records support batch review, investigations, traceability, and regulatory compliance.
15. Conclusion
Quality checks during pharmaceutical packing operations are not limited to inspecting the finished blister or carton. They begin with line clearance and material verification, continue through primary and secondary packaging, and conclude with reconciliation, documentation review, and controlled line closure.
A well-controlled packaging operation depends on three things: the right material, the right checks, and reliable documentation.
Production teams must understand the defects they are expected to identify, recognize when a process is no longer under control, and escalate discrepancies without delay. IPQC and QA must ensure that inspection results are evaluated appropriately and that deviations receive a documented, evidence-based assessment.
For pharmaceutical manufacturers, effective packing quality control protects product identity, labeling accuracy, traceability, and ultimately patient safety. A packing line should be considered successful not simply because it achieves its production target, but because it produces the required quantity of correctly packed, compliant, and traceable medicines.
Explore more practical GMP and pharmaceutical manufacturing guides on Pharma Manufacturing Hub
About the Author
Ramesh Palav is a pharmaceutical manufacturing professional with over 20 years of experience in pharmaceutical operations, oral solid dosage manufacturing, and GMP compliance. His areas of professional experience include tablet manufacturing, process and equipment qualification, validation, quality management systems, deviation management, CAPA, and regulatory audit readiness.
Through Pharma Manufacturing Hub, he shares practical knowledge, technical guidance, and industry-focused educational content to help pharmaceutical professionals strengthen their understanding of manufacturing operations, quality systems, regulatory compliance, and continuous improvement.
Website: https://pharmamanufacturinghub.com/